Articles

Nitrosamines in medicines: what they are, EMA and FDA limits and sources

13 min read

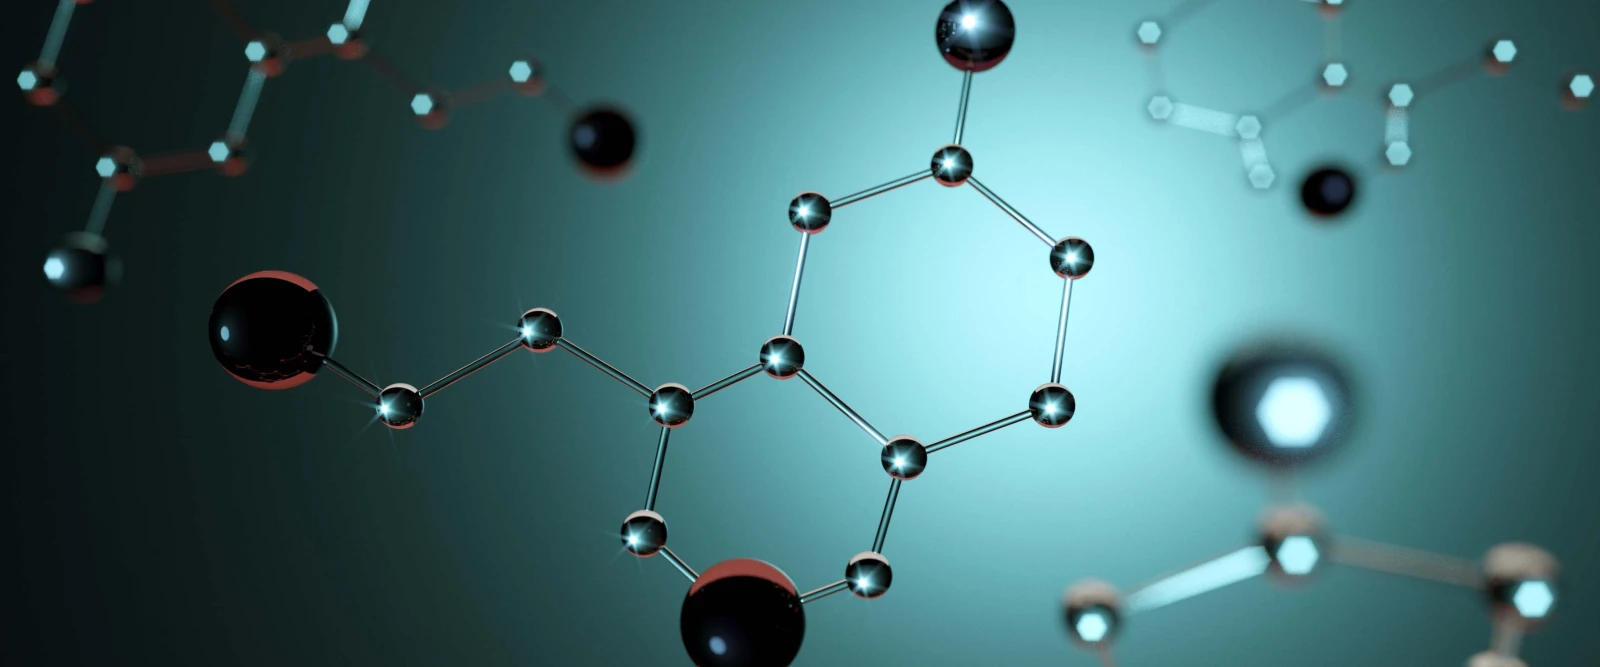
Nitrosamines (N-nitrosamines) are compounds in which a nitroso group is bonded to an amine nitrogen (R1R2N–N=O). Most are probable human carcinogens, so ICH M7 places them in the “cohort of concern” and regulators give each one its own acceptable intake: 96 ng/day for NDMA and 26.5 ng/day for NDEA, under both the EMA and the FDA.

Figures checked on 6 November 2026 against the EMA questions and answers for marketing authorisation holders (Rev. 24, 3 November 2026), the EMA list of acceptable intakes (Appendix 1, Rev. 14) and the FDA acceptable intake tables (updated 24 September 2026).

The N-nitroso group, R1–N–N=O, above tablets, capsules and cosmetics

What are nitrosamines?

They form when an amine — secondary, tertiary or quaternary — reacts with a nitrosating agent, typically nitrous acid produced from nitrites under acidic conditions. They have no industrial use of their own: they turn up as by-products in processed meat, alcoholic drinks, cosmetics, drinking water disinfected with chloramines and, since 2018, in medicines.

That year, N-nitrosodimethylamine (NDMA) was found in sartans, a class of blood-pressure medicines. The finding led the EMA to review all human medicines under an Article 5(3) referral, concluded in June 2020, and the FDA to publish its guidance on controlling nitrosamine impurities, now in its second revision (September 2024). Both require manufacturers to assess every product, test it where a risk exists and change the process or the specification when needed.

Most N-nitrosamines are classified by IARC as probably (group 2A) or possibly (group 2B) carcinogenic to humans. ICH M7(R2) puts N-nitroso compounds in the “cohort of concern”: mutagenic carcinogens so potent that the generic threshold of toxicological concern, 1.5 µg/day, cannot be applied to them by default. Each nitrosamine therefore gets its own acceptable intake (AI): the daily amount associated with roughly one additional cancer case in 100,000 people exposed every day for 70 years.

Small nitrosamines and NDSRIs

Regulators split nitrosamine impurities into two families:

  • Small-molecule nitrosamines do not resemble the active substance and turn up in many different products: NDMA, NDEA, NMPA, NDIPA, NIPEA, NDBA and NMBA.
  • Nitrosamine drug substance-related impurities (NDSRIs) carry the active substance, or a fragment of it, in their structure, so each one is specific to its API. They form mostly in the finished product, when the API reacts with nitrite left in the excipients. The EMA also calls them NO-API.

The difference matters because of the data behind the limit: the common small nitrosamines have carcinogenicity studies; most NDSRIs have none, so their limit comes from their structure (see CPCA below).

Acceptable intakes: EMA and FDA side by side

Lifetime acceptable intakes, in nanograms per day, for the small-molecule nitrosamines most often found in medicines:

NitrosamineCASEMA AI (ng/day)FDA AI (ng/day)
NDMA (N-nitrosodimethylamine)62-75-99696
NDEA (N-nitrosodiethylamine)55-18-526.526.5
NMBA (N-nitroso-N-methyl-4-aminobutyric acid)61445-55-41,500 (CPCA 4)1,500 (CPCA 4)
NMPA (N-nitroso-N-methylaniline)614-00-6100 (CPCA 2)100 (CPCA 2)
NEIPA / NIPEA (N-nitrosoethylisopropylamine)16339-04-1400 (CPCA 3)400 (CPCA 3)
NDIPA (N-nitrosodiisopropylamine)601-77-41,500 (CPCA 5)1,500 (CPCA 5)
NDBA (N-nitrosodi-n-butylamine)924-16-326.5—
NMOR (N-nitrosomorpholine)59-89-2127—
NPIP (N-nitrosopiperidine)100-75-41,300—
NPYR (N-nitrosopyrrolidine)930-55-21,700—
NMPEA (N-nitroso-N-methyl-2-phenylethylamine)13256-11-68—

—: no compound-specific value in the FDA tables as of 24 September 2026. Sources: EMA, Appendix 1, Rev. 14; FDA, CDER Nitrosamine Impurity Acceptable Intake Limits, tables 1 and 2.

The two agencies agree on all six small nitrosamines the FDA lists. The EMA list is far longer — 256 N-nitrosamines, most of them NDSRIs — and includes the lowest value of all, 8 ng/day for NMPEA. When no limit can be derived by any of its approaches, the FDA recommends 26.5 ng/day.

CPCA: limits for NDSRIs without data

The Carcinogenic Potency Categorisation Approach (CPCA) assigns an NDSRI to one of five potency categories from its structure alone. It rests on α-hydroxylation — the metabolic activation of the carbon next to the N-nitroso group — and scores the structural features that make it easier or harder.

CPCA categoryEMA AI (ng/day)FDA AI (ng/day)
11826.5
2100100
3400400
41,5001,500
51,5001,500

Category 1 is the only difference: 18 ng/day is the EMA’s class-specific threshold for N-nitrosamines, while the FDA uses 26.5 ng/day, based on NDEA, for products marketed in the United States. Category 2 reflects NDMA and NNK; categories 4 and 5 take the ICH M7 threshold of 1,500 ng/day.

How the category is reached (EMA, Appendix 2; FDA, NDSRI guidance, appendix A):

  1. Straight to category 5 if the molecule has no hydrogen on its α-carbons, if neither side of the N-nitroso group has more than one α-hydrogen, or if one α-carbon is tertiary.
  2. Potency score for the rest: an α-hydrogen score of 1 to 3, depending on how many hydrogens each α-carbon carries; plus points for deactivating features, such as a carboxylic acid (+3), a pyrrolidine ring (+3), a five- or six-membered ring (+2) or a morpholine (+1); minus one point for each activating feature, such as an aryl group on an α-carbon or a methyl group on a β-carbon.
  3. Score to category: 1 or less, category 1; 2, category 2; 3, category 3; 4 or more, category 4.

Worked example: NMBA scores 1 for its α-hydrogens and 3 for its carboxylic acid. Total 4: category 4, 1,500 ng/day, the category both agencies assign to it. The CPCA does not apply to N-nitrosamides, nitrosoureas, nitrosoguanidines or N-nitroso groups inside a heteroaromatic ring.

From ng/day to ppm: the specification limit

The AI becomes a specification limit when it is divided by the maximum daily dose on the label:

Limit (ppm) = AI (ng/day) ÷ maximum daily dose (mg/day)

The EMA’s own example, for a product taken at up to 300 mg a day: NDEA, 26.5 ÷ 300 = 0.088 ppm (88 ppb); NDMA, 96 ÷ 300 = 0.32 ppm (320 ppb). The calculation ignores the molecular weight of the nitrosamine, and the same AI applies to every route of administration.

Three rules decide what goes into the specification:

  • The 10% threshold. For the EMA, nitrosamines below 10% of their AI are a negligible risk. The FDA does not require a specification below that level either, provided the root cause is understood and the process is validated.
  • Limit of quantification (EMA, question 9): at or below the limit for routine testing, at or below 30% of it to justify skip testing, and at or below 10% of it to leave the nitrosamine out of the specification.
  • Nitrosamines from an excipient (new in Rev. 24): the limit is calculated on the maximum daily amount of that excipient at the maximum dose.

More than one nitrosamine

By default, both agencies cap the total at the AI of the most potent one. Both also accept a flexible approach in which each nitrosamine is limited to a percentage of its own AI and the percentages add up to 100% at most. The EMA frames it as a total risk no higher than 1 in 100,000; the FDA allows it for up to three nitrosamines and asks to be consulted beyond that.

Short treatments: less-than-lifetime limits

Since autumn 2026, both agencies allow the AI to be multiplied when the treatment lasts less than a lifetime:

Treatment durationEMA (Q&A Rev. 24, November 2026)FDA (24 September 2026)
Up to 1 month80 × AI80 × AI
More than 1 and up to 12 months13.3 × AI13.3 × AI
More than 1 and up to 10 years6.7 × AI6.67 × AI
More than 10 years to lifetimeNo adjustmentNo adjustment

The FDA’s own example: NDEA in a treatment of under 30 days, 26.5 × 80 = 2,120 ng/day. The duration has to match the labelling. The FDA does not apply the adjustment to interim limits, and for an approved product a higher AI is usually a major change.

Deadlines and where things stand in 2026

Both agencies set the same three steps: risk evaluation, confirmatory testing and changes to the marketing authorisation.

Risk evaluationConfirmatory testingChanges submitted
EMA, chemically synthesised APIs31 March 202126 September 20221 October 2023
EMA, biological APIs1 July 20211 July 20231 July 2023
FDA, small-molecule nitrosamines31 March 2021When a risk is identified1 October 2023
FDA, NDSRIs1 November 2023When a risk is identified1 August 2025
  • EMA. The deadlines have passed, but the reporting obligations still apply. Since November 2026, holders can apply interim limits for up to seven years from the publication of the first AI while they implement corrective actions, under the conditions of question 22.
  • FDA. On 23 June 2025 the FDA accepted that 1 August 2025 was not achievable for every application, asked for a progress report by that date and later extended the interim NDSRI limits to 1 August 2027. As of 24 September 2026 it had not published a new calendar.

Where nitrosamines enter a pharmaceutical process

The EMA (question 4) and the FDA list the root causes found since 2018. Beyond the chemistry of the API, several have to do with water, equipment and materials:

  • Synthesis. Nitrites or other nitrosating agents in contact with secondary or tertiary amines. Solvents such as DMF, DMAc or NMP can bring the amine.
  • Excipients. The FDA notes that many contain nitrite at ppm levels, the usual route to NDSRIs.
  • Water. In one documented case, chloramine used to disinfect the incoming water reacted with dimethylamine leaching from an anion-exchange resin, and NDMA ended up in water for injection.
  • Equipment and cross-contamination. Different processes run one after another on the same line, recovery processes in non-dedicated equipment and inadequate cleaning between products. Fluid-bed drying at high temperature and jet milling can also nitrosate amines with nitrogen oxides from the air.
  • Packaging. Heat-sealing blisters with nitrocellulose lidding releases nitrogen oxides. Rev. 24 adds polymer bags for parenteral products, where monomers, plastic additives and printing inks can bring amines and nitrosating agents. In August 2025 the FDA published information on NDBA and other small nitrosamines leaching from infusion bags.

The FDA states it plainly: container closure systems, secondary packaging included, and manufacturing equipment can be a source of nitrite or nitrosamines, and that risk belongs in extractables and leachables studies.

Rubber, elastomers and vulcanisation accelerators

The link between rubber and nitrosamines has been documented since the early 1980s. Rubber chemicals made from secondary amines, such as thiurams, dithiocarbamates and some sulfenamide accelerators, contain nitrosamines, and the rubber articles made with them contain the nitrosamines that correspond to those amines, especially when nitrosating agents such as nitrogen oxides are present during manufacture (Spiegelhalder and Preussmann, 1982 and 1983).

The rule is simple: the nitrosamine follows the amine of the accelerator.

Accelerator derived fromExampleNitrosamine
DimethylamineTetramethylthiuramNDMA
DiethylamineZinc diethyldithiocarbamate (ZDEC)NDEA
DibutylamineZinc dibutyldithiocarbamateNDBA
MorpholineN-oxydiethylene benzothiazole sulfenamideNMOR

The pairs apply the correspondence rule described in the sources; the ZDEC to NDEA pathway is the one drawn explicitly, by the German BfR in 2002.

That BfR opinion, on balloons, found NDMA, NDEA and NDBA and recommended accelerators based on “safe” amines that are hard to nitrosate. In 1993, Directive 93/11/EEC set release limits for rubber teats and soothers: 0.01 mg/kg of N-nitrosamines and 0.1 mg/kg of nitrosatable substances.

For medicines, the EMA’s 2020 assessment report lists emissions from vulcanisation among the theoretically possible causes, including rubber in contact with the product. The Q&A also warns of the opposite problem: traces of nitrosamines in rubber or plastic labware can give false positives in the analysis.

No regulator classifies a whole family of materials as nitrosamine-free. What carries weight is data on the specific compound: its curing system, its raw materials and its extractables.

What to ask the supplier of a product-contact component

The EMA expects manufacturers and their raw-material suppliers to give marketing authorisation holders everything needed for a complete risk evaluation (question 7), and changes of supplier, process or packaging to be controlled because they can change the risk (question 12). The FDA asks for container closures and equipment to be covered by extractables and leachables studies, and for suppliers to be qualified under ICH Q10.

No agency publishes a questionnaire. Based on the above, three questions are worth putting to the supplier of a hose, a tube, a gasket or a single-use assembly:

  1. Which curing system does the compound use, and does it contain accelerators derived from secondary amines, such as thiurams, dithiocarbamates or sulfenamides?
  2. Do the extractables and leachables data include nitrosamines (at least NDMA, NDEA, NDBA, NMOR, NPIP and NPYR) and their precursor amines?
  3. Will the supplier notify changes of formulation or of raw-material supplier?

For Venair’s silicone products, the position is the one the company has published since 2023: “We exclusively use nitrosamine and nitrosatable-free raw materials sourced from trusted and accredited suppliers and we employ qualified production processes that ensure that none of our silicone-based products incorporate hazardous substances.”

Venair’s sanitary and pharmaceutical hoses and single-use tubing and bioprocess assemblies are described in the Life Science 2027 catalogue. To check how a material holds up against a cleaning agent or a solvent, see the chemical compatibility chart. On biocompatibility, read what USP Class VI tubing is; on substances of very high concern, the REACH regulation.

Frequently Asked Questions

What are nitrosamines?

Nitrosamines, or N-nitrosamines, are compounds with a nitroso group bonded to an amine nitrogen (R2N–N=O). They form when amines react with nitrosating agents such as nitrites under acidic conditions, and they appear as by-products in food, drinking water, cosmetics and medicines.

Why are nitrosamines dangerous?

Most are mutagenic and probably or possibly carcinogenic to humans (IARC groups 2A and 2B). ICH M7 places them in the “cohort of concern”, so each one gets its own acceptable intake instead of the generic threshold of 1.5 µg/day.

What is the acceptable intake of NDMA?

96 ng/day, for both the EMA and the FDA. For a medicine taken at up to 300 mg a day, that is a limit of 0.32 ppm.

What is an NDSRI?

A nitrosamine drug substance-related impurity: a nitrosamine that contains the active substance, or a fragment of it, in its structure. It usually forms in the finished product when the API reacts with nitrite from the excipients, and when there are no carcinogenicity data its limit comes from the CPCA.

How is a nitrosamine limit calculated in ppm?

Divide the acceptable intake in ng/day by the maximum daily dose in mg/day. NDEA (26.5 ng/day) in a product taken at up to 300 mg a day gives 0.088 ppm.

Can rubber components release nitrosamines?

Yes, when the rubber is vulcanised with accelerators derived from secondary amines, such as thiurams and dithiocarbamates. The EMA lists vulcanisation emissions among the possible causes in medicines, and the FDA asks for packaging and equipment to be assessed in extractables and leachables studies.

References

  1. EMA. Questions and answers for marketing authorisation holders/applicants on the CHMP Opinion for the Article 5(3) referral on nitrosamine impurities in human medicinal products. EMA/409815/2020 Rev. 24, 3 November 2026.
  2. EMA. Appendix 1: Acceptable intakes established for N-nitrosamines. Rev. 14, October 2026.
  3. EMA. Appendix 2: Carcinogenic Potency Categorisation Approach for N-nitrosamines. EMA/451665/2023, 12 October 2023.
  4. EMA, CHMP. Assessment report. Nitrosamine impurities in human medicinal products, Article 5(3) procedure. EMA/369136/2020, 25 June 2020.
  5. FDA, CDER. Control of Nitrosamine Impurities in Human Drugs. Guidance for Industry. Rev. 2, September 2024.
  6. FDA, CDER. Recommended Acceptable Intake Limits for Nitrosamine Drug Substance-Related Impurities (NDSRIs). Guidance for Industry. August 2023.
  7. FDA, CDER. CDER Nitrosamine Impurity Acceptable Intake Limits. Updated 24 September 2026.
  8. FDA, CDER. Application of Less-Than-Lifetime Adjustment to Nitrosamine Impurities. 24 September 2026.
  9. FDA, CDER. Emerging Scientific and Technical Information on Leachable NDBA and Other Small-Molecule Nitrosamines in Infusion Bags. 18 August 2025.
  10. ICH. M7(R2) Assessment and Control of DNA Reactive (Mutagenic) Impurities in Pharmaceuticals to Limit Potential Carcinogenic Risk. Step 4, 3 April 2023.
  11. Commission Directive 93/11/EEC of 15 March 1993 concerning the release of the N-nitrosamines and N-nitrosatable substances from elastomer or rubber teats and soothers. OJ L 93, 17 April 1993.
  12. BfR (formerly BgVV). Risk assessment of N-nitrosamines in balloons. 11 April 2002.
  13. Spiegelhalder B., Preussmann R. Nitrosamines and rubber. IARC Scientific Publications, 1982 (41), 231–243.
  14. Spiegelhalder B., Preussmann R. Occupational nitrosamine exposure. 1. Rubber and tyre industry. Carcinogenesis, 1983, 4 (9), 1147–1152.
  15. Sung J.H. et al. (2010). Liquid chromatography tandem mass spectrometry determination of N-nitrosamines released from rubber or elastomer teats and soothers. Food Additives and Contaminants: Part A, 27 (12), 1745–1754.
  16. Kühne F. et al. (2018). N-nitrosamines migration from food contact materials into food simulants: analysis and quantification by means of HPLC-APCI-MS/MS. Food Additives and Contaminants: Part A, 35 (4), 792–805.
  17. Monteiro M.A. et al. (2023). Investigation of carcinogenic impurities of N-nitrosamines in sartan pharmaceutical products marketed in Brazil. Journal of Pharmaceutical Sciences, 112 (5), 1305–1314.

Related

All